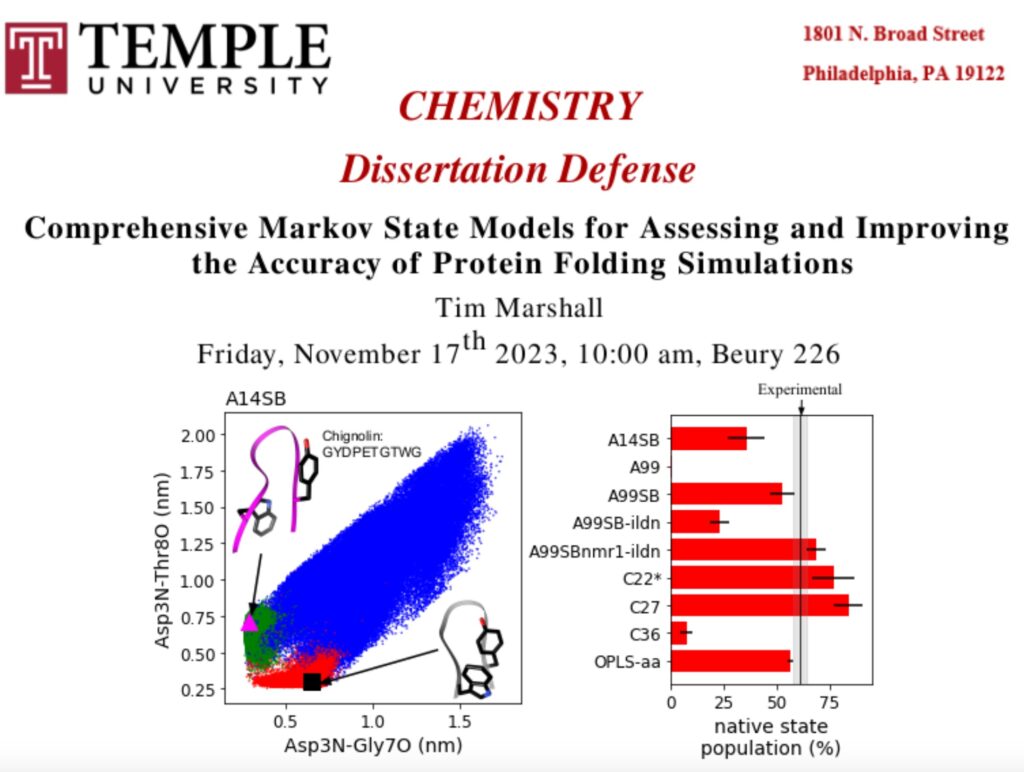
On Friday, November 17, Tim Marshall successfully defended his Ph.D Thesis “Comprehensive Markov State Models for Assessing and Improving the Accuracy of Protein Folding Simulations”.
Tim’s work has focused on using MSMs as a platform for evaluating force fields and for examining the ability of molecular simulations to quantitatively predict how mutations affect folding stability, toward the prospect of simulation-based virtual screening for protein design. Tim is still preparing several manuscripts describing this work, so stay tuned for preprints.
Model selection using replica averaging with Bayesian inference of conformational populations. Raddi RM, Marshall T, Ge Y, Voelz V. https://doi.org/10.26434/chemrxiv-2023-396mm
In this work, we describe and demonstrate with examples how equipping our BICePs algorithm with a replica-averaged forward model makes it a Maximum-Entropy reweighting method with some key advantages:
It is a post-processing method that does not require restraints implemented inside simulations
Heuristic estimates of uncertainty are not needed; posterior sampling enables learning uncertainties from the data
BICePs has improved likelihood models to account for outliers
The BICePs score can be used for objective model selection
Automated force field and forward model parameterization can be performed using the BICePs score as an objective function.
I recently spoke about this work at the ACS Fall 2023 symposium “Emerging Techniques to Quantify Biomolecular Conformational Ensembles” symposium.
Raddi, Robert M., Yunhui Ge, and Vincent A. Voelz. “BICePs v2.0: Software for Ensemble Reweighting Using Bayesian Inference of Conformational Populations.” Journal of Chemical Information and Modeling 63, no. 8 (April 24, 2023): 2370–81. https://doi.org/10.1021/acs.jcim.2c01296.
Congrats to Rob Raddi on this paper and for coding this more user-friendly and extensible Python package: https://github.com/vvoelz/biceps
With five students presenting at BPS, the Voelz Lab made a strong showing at BPS 2023 in San Diego, CA, Feb 18-22! Dylan Novack gave an early-morning (yet still riveting 🙂 Platform Session talk on work simulating FOXO1 and the effects of mutations. Here are the presentations:
An Evaluation of Force Field Accuracy for the Mini-Protein Chignolin using Markov State Models Tim Marshall, Robert Raddi, Vincent A. Voelz
Model selection using Bayesian inference of conformational populations with replica averaging Robert M. Raddi, Tim Marshall, Yunhui Ge, and Vincent A. Voelz
ONCOGENIC MUTATIONS IN THE DNA-BINDING DOMAIN OF FOXO1 THAT DISRUPT FOLDING: QUANTITATIVE INSIGHTS FROM EXPERIMENTS AND MOLECULAR SIMULATION. Dylan Novack Platform Session Title: Platform: Protein Stability, Folding, and Chaperones
De novo design of beta hairpin peptides with custom installed halogen-bonding groups Thi Dung Nguyen, Vincent Voelz
Computational fragment screening of dimeric SARS-CoV-2 Main Protease Rashad Reid Jr., Vincent A. Voelz
Last day of the conference, still rocking our lanyardsSpecial guest appearance by Voelz Lab alum George Pantelopulos!
Congratulations to Ph.D student Rob Raddi, a 2022 winner of the College of Science and Technology Outstanding Teaching Assistant (TA) at Temple University! This prestigious college-level award attests to Rob’s excellence in teaching. Great job and well deserved!
Our new paper “Multi-ensemble Markov Models to estimate affinities and rates of ligand binding is now published in the Journal of Chemical Physics. Congrats to Yunhui Ge for getting this across the finish line!
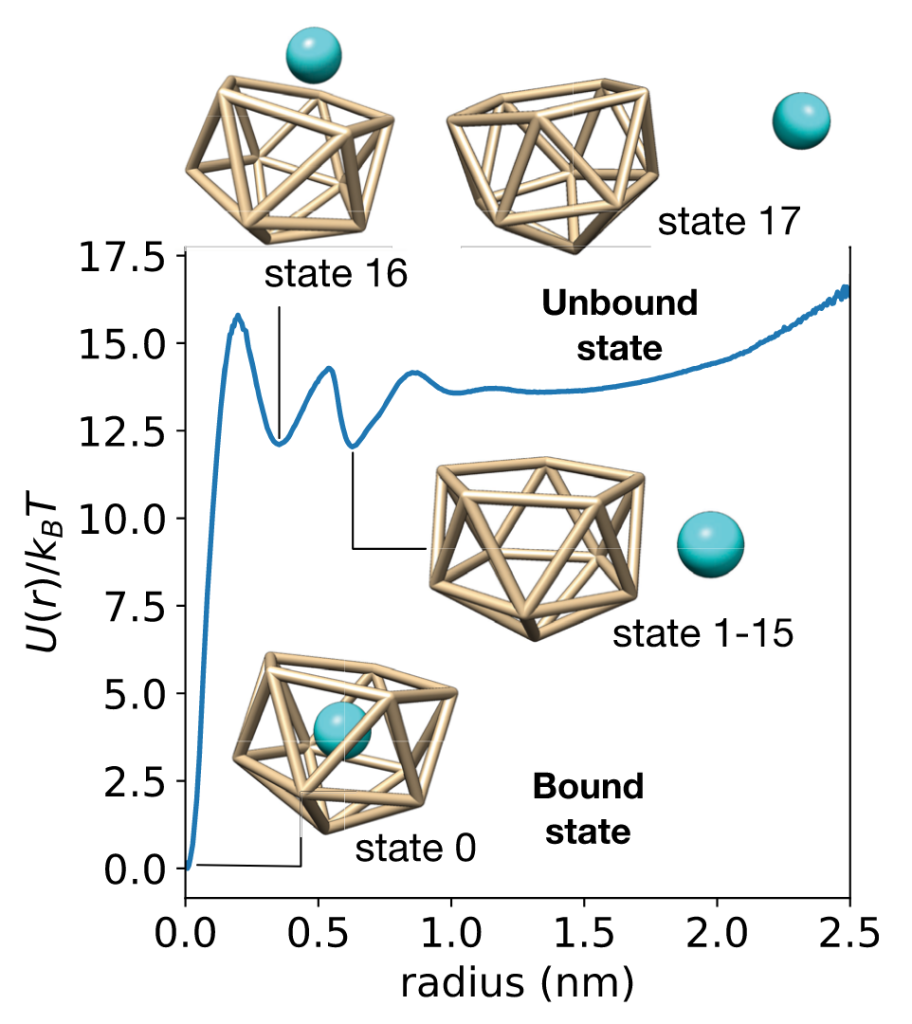
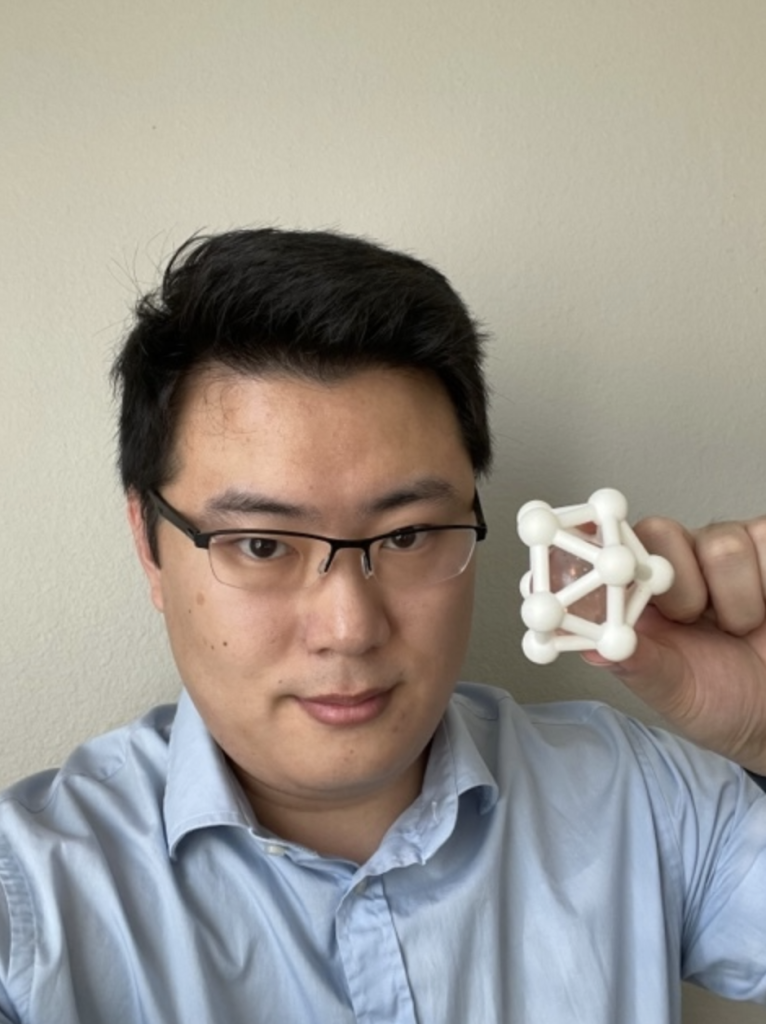
This work examines a toy receptor made from an icosahedron of 11 Lennard-Jones particles. The ligand is a LJ particle, and all is solvated in explicit water. This system has number of encounter-complex states and a slow(ish) residence time of 30.3 ns.
While conventional MSMs accurately estimate binding rates from swarms of short trajectories, they estimate affinities poorly. Why? Because MSM estimators typically enforce detailed balance, assuming the data is sampled at equilibrium.
So even with lots of parallel simulation, it still takes a long time for bound and unbound populations to equilibrate, and get a good model. (Our related work explored this problem with adaptive seeding simulations.)
Instead, what if we could quickly collect many binding/unbinding transitions in a biased ensemble, and use this information to infer populations and rates in the unbiased ensemble. This is a job for multiensemble Markov models, or MEMMs!
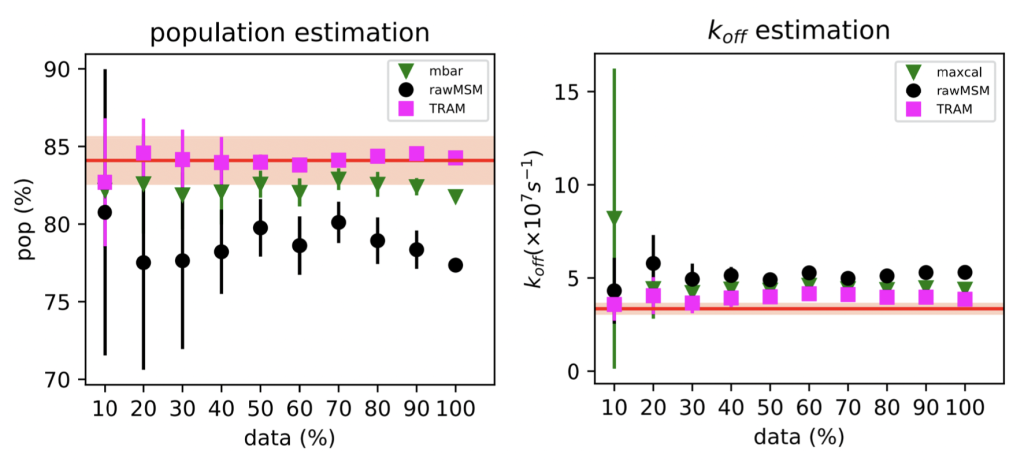
To accelerate transitions, we scaled the ligand nonbonded interactions, as is typical in FEP calculations for drug discovery. We tried two different MEMM estimators: (1) TRAM, from the Frank Noé group in Berlin, and (2) our Maximum-Caliber approach.
We find that both work pretty well (!), but TRAM works slightly better.
These results have encouraged us to explore how this method works in more realistic ligand binding systems, as a potential tool for MD-based virtual screening. Expect more from our lab on this soon 🙂
You can read more about this work here:
Estimation of binding rates and affinities from multiensemble Markov models and ligand decoupling. Ge, Yunhui, and Vincent A. Voelz. The Journal of Chemical Physics 156, no. 13 (April 7, 2022): 134115. https://doi.org/10.1063/5.0088024
Yunhui Ge, holding a model of the toy binding pocket.
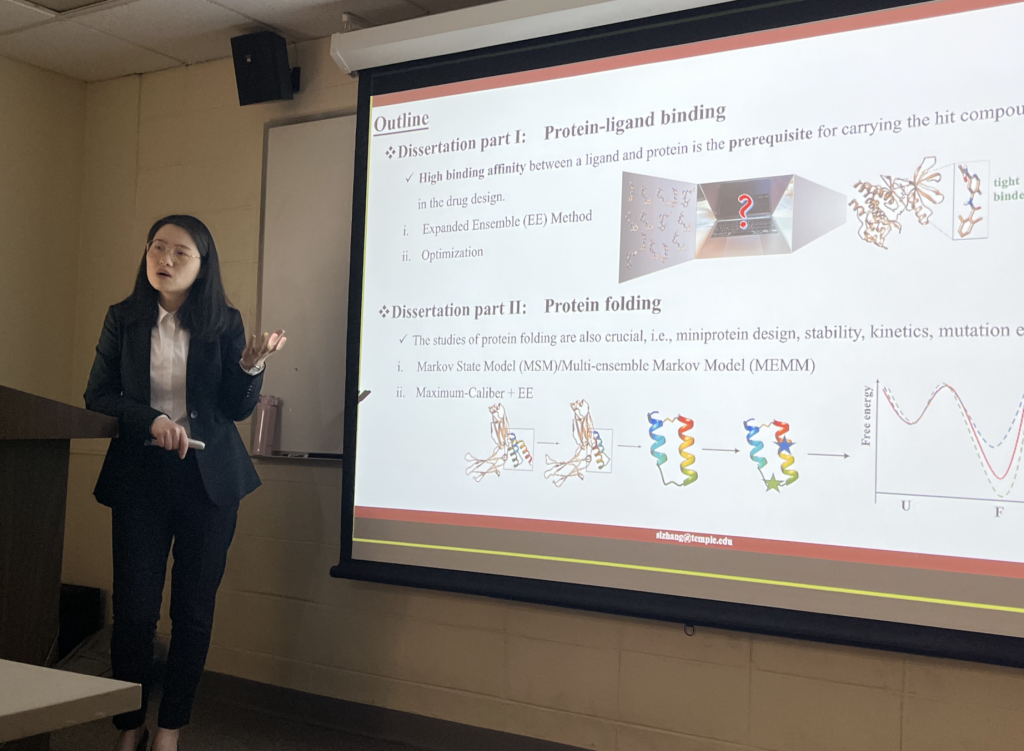
On April 5, Si Zhang successfully defended her PhD thesis “Computational Approaches for Protein Folding and Ligand Binding: from Thermodynamics to Kinetics”. Congratulations Si!
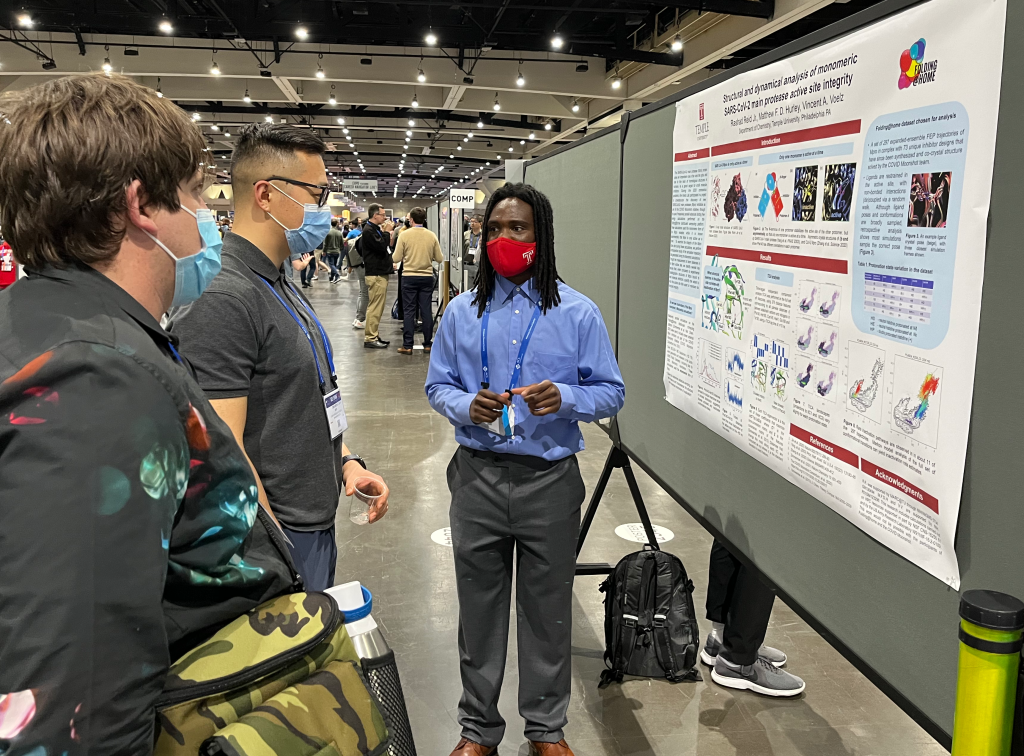
Congrats to undergrad researcher and MARC scholar Rashad Reid, who presented his poster “Structural and dynamical analysis of monomeric SARS-CoV-2 Mpro active site integrity” at the Spring 2022 Meeting of the American Chemical Society in San Diego this week.
Rashad was selected for both the ACS COMP session and the SCI-MIX. Great work, Rashad!